• Skip to primary navigation
  • Skip to main content
  • Skip to footer
ElevateBio

ElevateBio

Powering the creation of cell & gene therapies at a speed the world deserves.

  • Resources
    • Resources Overview
    • Elevated Insights
    • Presentations & Publications
    • Whitepapers
  • Media
    • Media Overview
    • Press Releases
    • Featured News
  • Facilities
  • Careers
  • Contact
  • Search
  • Manufacturing & Discovery Services
    • Manufacturing & Discovery Services Overview
    • Gene Editing Design & Optimization
    • Process Development
    • Analytical Development
    • cGMP Manufacturing
    • Next-Generation Sequencing
    • Quality
    • Regulatory CMC
    • Certifications & Compliance
    • Generative AI
  • Technologies
    • Technologies Overview
    • Gene Editing
    • Viral and Non-Viral Delivery
    • Messenger RNA (mRNA)
    • Induced Pluripotent Stem Cells (iPSCs)
  • Advanced Therapies
  • Partner With Us
  • About
    • About Overview
    • Our Journey
    • Our Team
    • Culture of Expedition
    • Careers
    • Investors
  • Resources
    • Resources Overview
    • Elevated Insights
    • Presentations & Publications
    • Whitepapers
  • Media
    • Media Overview
    • Press Releases
    • Featured News
  • Facilities
  • Careers
  • Contact
  • Manufacturing & Discovery Services

    Manufacturing & Discovery Services Overview

    Capabilities & Services
    • Gene Editing Design & Optimization
    • Process Development
    • Analytical Development
    • cGMP Manufacturing
    • Next-Generation Sequencing
    Expertise
    • Quality
    • Regulatory CMC
    • Certifications & Compliance
    • Generative AI
  • Technologies
    • Technologies Overview
    • Gene Editing
    • Viral and Non-Viral Delivery
    • Messenger RNA (mRNA)
    • Induced Pluripotent Stem Cells (iPSCs)
  • Advanced Therapies
  • Partner With Us
  • About
    • About Overview
    • Our Journey
    • Our Team
    • Culture of Expedition
    • Careers
    • Investors

Manufacturing

September 3, 2026 by

The future of advanced therapies won’t be defined by a single technology. For years, our industry has searched for the next breakthrough, whether that’s a more precise gene editor, a more efficient manufacturing platform, a faster analytical method, or an emerging modality poised to transform medicine. Each innovation matters, but none alone is enough to address the complexity of developing the next generation of therapies.

Too often, these technologies have evolved independently, with discovery, manufacturing, analytics, and emerging modalities advancing along separate paths. Enabling the next generation of therapies will require continuous investment across the entire advanced therapy lifecycle, from therapeutic discovery and process development to manufacturing, quality, release, and the technologies that will define the future of medicine.

At ElevateBio, that’s the philosophy driving our technology strategy. We are continuously investing in the capabilities needed to advance therapies from discovery through commercialization, because meaningful innovation happens when every stage of the lifecycle is strengthened. Every investment we make is intended to solve today’s challenges while building the infrastructure needed for tomorrow’s therapies.

Gene Editing Requires Multiple Approaches

Gene editing has transformed our understanding of what is therapeutically possible. That’s why ElevateBio built a diversified gene editing portfolio that includes novel nucleases and base editors, programmable gene insertion technologies, epigenetic editors, AI-designed enzymes, and the manufacturing capabilities needed to translate these discoveries into clinical development. This breadth allows us to approach complex disease targets with greater flexibility while reducing development risk and increasing the likelihood of identifying the optimal therapeutic strategy.

AI is accelerating this evolution even further. By combining a proprietary library of more than 10 billion proteins with generative AI and machine learning, we’re expanding protein engineering beyond the limits of naturally occurring biology. Rather than relying solely on existing proteins, we can computationally design and optimize therapeutic candidates with improved precision, potency, and safety. As AI continues to reshape biotechnology, its greatest impact may not simply be making discoveries faster, but helping scientists explore entirely new possibilities in biology that were previously inaccessible.

Together, these investments represent a continued commitment to expanding the possibilities of gene editing. By building capabilities across multiple technologies, platforms, and approaches, we are creating the foundation to address increasingly complex therapeutic challenges.

Automation Is Redefining Cell Therapy Manufacturing

Scientific innovation only creates value if it can be manufactured consistently and at scale. Cell therapy manufacturing remains one of the industry’s greatest operational challenges. Manual workflows can cause variability, limit scalability, and contribute significantly to costs. Addressing these challenges requires more than incremental process improvement; it requires rethinking how manufacturing itself is designed.

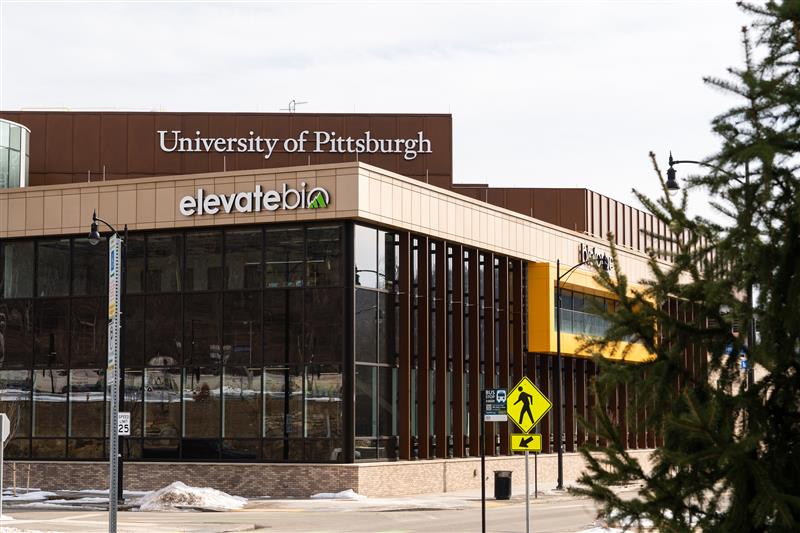
We recently completed building our new advanced therapies manufacturing facility in Pittsburgh and expect it to be operational next year. At this facility, we are taking a fresh look at automating manufacturing to reduce complexity, improve scalability, and lower cost of goods sold. This includes fully integrating and leveraging all the data and process systems we utilize to minimize human error, improve process consistency, accelerate production timelines, and create a foundation for more efficient manufacturing. We’ve also collaborated with companies such as Sartorius and OriBiotech on the optimization and implementation of cell therapy platforms. The objective is not simply to replace manual steps with automated ones, but to leverage systems that are inherently more predictable, reproducible, and efficient.

As advanced therapies move toward broader commercialization, manufacturing excellence will become just as important as scientific excellence.

Data Should Accelerate Decisions, Not Delay Them

Innovation doesn’t stop once manufacturing is complete. Release testing remains one of the most time-intensive stages of advanced therapy manufacturing, and every additional day spent waiting for analytical results delays treatment and adds operational complexity. We see an opportunity to fundamentally change this process by making analytics a more integrated, data-driven, and efficient part of the manufacturing workflow.

By investing in developing next-generation sequencing (NGS), rapid analytics, automation, and connected data workflows, we’re building release testing capabilities that compress timelines while maintaining the rigorous analytical standards these therapies require. Traditional release can often take weeks, representing upwards of 50% of overall manufacturing time. Whereas at ElevateBio, new sequencing assays can deliver results in as little as four days, helping reduce a critical bottleneck and accelerate the path to release. Our vision extends beyond implementing new instruments. We’re creating integrated analytical workflows where sequencing, data analysis, interpretation, and reporting work together to enable faster release decisions, reduce testing complexity, and improve manufacturing efficiency.

In the future, analytics won’t simply verify quality; they will become a driver of manufacturing innovation. The data generated throughout manufacturing and release can provide valuable insights to optimize processes, strengthen platform capabilities, and continuously improve future therapies.

Investing Ahead of the Next Wave of Innovation

Perhaps the most important technology investments are the ones the market hasn’t fully demanded yet. Driving innovation requires looking beyond today’s challenges and building the capabilities that will enable tomorrow’s therapies. The field continues to evolve rapidly, with mRNA and lipid nanoparticle (LNP) technologies emerging as foundational tools that extend well beyond vaccines, including in vivo cell therapies.

Recognizing that shift, we’ve continued investing in building capabilities spanning our lipid library, formulation, process development, quality systems, and manufacturing for mRNA and LNP-based programs. We have already manufactured over 25 drug substance or bulk drug product mRNA batches for both vaccine and in vivo therapeutic applications. We have also developed our own end-to-end mRNA manufacturing process, which we anticipate making available to partners by early 2027. Just as importantly, we’re building flexibility into our manufacturing model. Some partners arrive with established manufacturing processes ready for GMP execution, while others need support much earlier in development. Our vision is to meet innovators wherever they are by providing an integrated path from development through commercial production. Reducing technical transitions ultimately reduces development risk, shortens timelines, and helps innovative therapies move more efficiently toward patients.

Advancing Innovation Across the Advanced Therapy Lifecycle

Advanced therapies are becoming increasingly sophisticated. The innovations that define the next decade will likely require continued investments across multiple technologies and capabilities, rather than standalone platforms, making flexibility itself a critical capability for enabling the future of medicine.

At ElevateBio, we believe driving innovation requires building capabilities today that can address the challenges of tomorrow. By continuously investing across the advanced therapy lifecycle, we are strengthening the tools and technologies that enable discovery and development, improving manufacturing scalability, accelerating decision-making, and preparing for the therapies of the future.
Mike Paglia, Chief Technology Officer

Michael Paglia is the Chief Technology Officer at ElevateBio, a technology-driven company commercializing its enabling technologies, manufacturing capabilities, and industry-leading expertise to accelerate the development of genetic medicines to treat human diseases. He has more than two decades of experience in facility design, start up, and operations ensuring the highest standards of quality, safety, and regulatory compliance. 

At ElevateBio, Michael led the design, construction and operations of the BaseCamp manufacturing facility that was recognized as the Facility of the Year, Operational Excellence by International Society for Pharmaceutical Engineering (ISPE) in 2021.  Michael established the process development and manufacturing capability and leads manufacturing operations, CMC regulatory, process/analytical development, and the advancement of innovative process technologies. 

Prior to ElevateBio, Michael was the Vice President of CMC Operations at Oncorus responsible for the development and manufacturing of novel genetically modified oncolytic herpes virus for the treatment of cancer and prior to that, Head of Technical Operations, Cellular Process Development and Manufacturing Operations at bluebird bio where he led the early process development, manufacturing, and global technology transfer of four approved genetically modified autologous cell therapies.  Early in his career at Tolerx, Michal lead process development, and late-stage manufacturing of novel therapeutic antibody products designed to treat patients by reprogramming the immune system.

Michael received his undergraduate degree from Providence College and a Master’s of Science in Biochemistry and Cellular Biology from the University of New Hampshire where he was honored with the Distinguished Alumni Award from the College of Life Science and Agriculture (COLSA) in 2023 for his career guidance and ongoing initiatives in COLSA to enhance STEM workforce development initiatives.
August 20, 2026 by

The best time to engage a cell and gene therapy CDMO is when key process development decisions are still being made, not once your process is ready for GMP manufacturing. Early manufacturing input reduces risk, accelerates technology transfer, and improves manufacturability and process consistency from the start.

Most Programs Engage Their CDMO Too Late

When we ask when the right time is to engage a CDMO, the answer we hear most often is, “Once the process is ready for technology transfer”. Nobody wants to pull a busy technical team into CDMO conversations before there’s a process ready to hand off. But by then, the decisions that shape manufacturability, scalability, and operational success are already locked in. A process that performs well in the lab can still hit real problems once it moves into GMP manufacturing.

Manufacturing Experience Matters More Than Capacity

Facilities, equipment, and people are the obvious assets a CDMO brings to the table. The more valuable one is manufacturing experience, the practical knowledge of how a process actually performs in a GMP setting. The right time to draw on that expertise is during process development, not after the process is locked, so the process gets designed with manufacturability in mind from the start.

A CDMO Sees Manufacturing Risks a Development Lab Can’t

A CDMO can tell you things a development environment can’t. Equipment reliability is one: how often does that piece of equipment generate errors when it’s run over and over, batch after batch? Raw materials and vendors are another: does the vendor provide robust quality control, and are they a supply partner you can actually count on? Timing matters too. An unstable intermediate that has to be prepared just in time can take longer to process on the GMP floor than it did on the lab bench. And in-process test results, the ones that trigger decisions like seeding, splitting, or harvesting, need a home: on the manufacturing floor or in the QC lab.

The Right CDMO Shapes Your Process, Not Just Manufactures It

The right question when evaluating a cell and gene therapy CDMO isn’t whether they can manufacture your process. It’s whether they have the process development capability to engage before the process is even defined. A CDMO brought in at the point of clinical manufacturing inherits your constraints. A partner engaged during process development helps you avoid creating them in the first place, so your program never loses momentum handing off from development to manufacturing.

ElevateBio Integrates Process Development and Manufacturing From the Start

ElevateBio offers a strong integrated tech transfer team that includes SMEs from manufacturing operations, MSAT, and process development. They provide critical manufacturability design input to the process development effort upfront, rather than a manufacturability assessment after the fact that points out what’s wrong with a process that’s already been transferred. This early engagement doesn’t just derisk and accelerate tech transfer. It also reduces variability, and failures in clinical manufacturing.

Yogesh Waghmare, Senior Director, MSAT

LinkedIn

Yogesh Waghmare, PhD is Senior Director of Manufacturing Science and Technology at ElevateBio, where he leads end-to-end technical support for GMP manufacturing of advanced modalities including cell therapies, viral vectors, and mRNA. With over 15 years of experience across MS&T, process development, and CMC leadership, he has held senior roles at Moderna, Editas Medicine, bluebird bio, and Sanofi-Genzyme, supporting clinical through commercial programs and regulatory submissions. Yogesh holds a PhD in Chemical Engineering and has authored numerous publications and delivered invited presentations in bioprocessing and advanced manufacturing.

August 20, 2026 by

In 2023, Kyverna Therapeutics and ElevateBio formed a partnership with a bold mission: to be the first to deliver cutting-edge CAR T-cell therapies to patients with autoimmune diseases. Today, that collaboration is yielding tangible results. Kyverna initiated a rolling Biologics License Application (BLA) submission for miv-cel (mivocabtagene autoleucel, KYV-101) for stiff person syndrome (SPS) in May 2026 – the first BLA submission for a CAR T-cell therapy for an autoimmune disease. This case study explores how Kyverna is leveraging ElevateBio’s process development and manufacturing capabilities to advance miv-cel towards commercialization.

The Challenge: Rapid Manufacturing Without Compromise

When Kyverna initially partnered with ElevateBio, the company needed to quickly execute a technology transfer of its autologous CAR T-cell therapy (miv-cel). But this was more than a handover of equipment. It started with documentation and required navigating compressed timelines, with a need to complete comparability quickly.

Any delay had the potential to impact clinical trials, patients, and regulatory process – which is why Kyverna needed a partner with the speed, flexibility, and commercial mindset to deliver a true blueprint for accelerated execution without sacrificing quality.

An Advanced Therapy CDMO Built for Acceleration

Unlike a traditional CDMO solely focused on transactional production services, ElevateBio helps biopharmaceutical companies design, develop, and manufacture therapies faster and more efficiently. With integrated discovery, process, and manufacturing capabilities, ElevateBio is redefining how advanced therapies are developed and scaled, acting as a true partner in innovation. Through its partnership, Kyverna received immediate access to ElevateBio’s end-to-end capabilities, expert teams in process development, analytical development, and manufacturing. Around the table, the two teams collaborated closely to advance towards a common mission of bringing potential life-transformative therapies to patients.

ElevateBio Accelerated Tech Transfer Without Compromising GMP Quality

The average industry technology transfer can take 12-18 months. When Kyverna began working with ElevateBio, they needed to move even faster to support an active IND filing while building a scalable foundation for future clinical and commercial development.

ElevateBio was able to complete the first comparability runs in the GMP environment within 4 months. Project kickoff to GMP readiness runs to support regulatory milestones and site qualification. Notably, the speed and rigor of this early collaboration established the operational and quality framework that helped support Kyverna’s advancement toward its recent BLA submission.

Key achievements included:

  • Developed and optimized new GMP-ready batch records, manufacturing documentation, and fill instructions to support rapid clinical manufacturing readiness and long-term scalability
  • Executed a successful comparability campaign with consistent right-first-time execution, minimizing runs, reducing risk, and keeping timelines intact
  • Leveraged in-house analytical and quality systems tightly connected to manufacturing – enabling faster coordination, rapid data turnaround, and streamlined execution without reliance on third-party testing
  • Evaluated and qualified analytical assays and microbiological safety methods ahead of comparability runs, helping ensure robust data generation and proactive risk mitigation
  • Completed all comparability successfully supporting accelerated data delivery for critical regulatory timelines and avoided delays to clinical development

Accelerating a full technology transfer while maintaining rigorous GMP and quality standards is a highly complex achievement – one that reflects the agility, cross-functional coordination, and technical expertise of the teams involved. The rapid transfer established a strong operational and regulatory foundation that helped support the continued advancement of miv-cel toward commercialization.

A Partnership Built for Scalability  

As Kyverna advanced miv-cel through clinical development and toward its rolling BLA submission, ElevateBio served as a strategic manufacturing partner supporting the program’s evolving operational and regulatory needs. Together, the teams advanced Kyverna’s miv-cel platform, helping support the speed, coordination, and quality execution required for a complex cell therapy program.

Key elements of the partnership included:

  • ElevateBio’s manufacturing capability enabled the speed, coordination, and continuity required for a complex, patient-specific cell therapy program, ensuring seamless execution across regions and timelines
  • ElevateBio’s commercial-ready infrastructure and quality systems provided confidence that manufacturing will scale alongside the program’s advancement. ElevateBio’s ICMC ™ certification for cell and gene therapy manufacturing further reinforced GMP excellence and future readiness
  • A flexible, highly collaborative operating model allowed both teams to rapidly adapt to evolving program needs, maintaining speed without compromising quality or execution standards

Together, the partnership demonstrates how integrated manufacturing expertise, operational flexibility, and aligned execution can help accelerate complex cell therapy programs from early development through regulatory milestones and toward commercialization readiness.

Building the Foundation for Regulatory Success

By combining accelerated technology transfer, integrated manufacturing execution, and close cross-functional collaboration, Kyverna and ElevateBio established a strong operational foundation that helped rapidly advance miv-cel through pivotal clinical development and key regulatory milestones, including BLA submission for SPS.

What began as an accelerated technology transfer evolved into a strong partnership spanning process development, GMP manufacturing, analytical support, and clinical supply execution. Across this continuum, the collaboration demonstrated that speed and rigor are not competing priorities, but rather mutually reinforcing priorities when enabled by integrated capabilities and aligned execution.

This enabled Kyverna to progress the first CAR T-cell therapy in autoimmune disease through a key regulatory milestone. Importantly, every milestone was guided by a single objective: delivering a transformative therapy to patients living with autoimmune disease.

In July 2026, ElevateBio announced a commercial manufacturing and supply agreement with Kyverna for both U.S. commercial and global clinical supply of miv-cel, extending our multi-year partnership.

July 9, 2026 by

ElevateBio is industrializing cell and gene therapy manufacturing to bring life-changing genetic medicines to improve the lives of patients and their families.

Henry Ford didn’t invent the automobile, but in 1913, he transformed how it was made. The moving assembly line drastically reduced the cost of a car and turned a luxury into something accessible to millions. Today, a parallel transformation is underway in medicine. Cell and gene therapies represent a fundamentally new class of treatment. Rather than managing symptoms with traditional drugs, these advanced therapies harness a patient’s own cells or genes to correct disease at its source, offering the potential for life-transformative therapies or even single-treatment cures. 

Since the first FDA-approved cell therapy in 2017, the field has moved from scientific breakthrough to medical reality. But manufacturing and delivering these therapies at scale remains the greatest challenge, and one that we at ElevateBio, a U.S.-based biomanufacturing company, are working to solve.

From Massachusetts to Pennsylvania: Building the Future of Medicine

The science of genetic medicine has reached an inflection point. Decades of research have yielded multiple approved cell therapies. Yet accessibility remains constrained, not by what’s biologically possible, but by how these medicines are manufactured. Cell therapy manufacturing is inherently personal: very often, one patient’s cells must be collected, engineered, and returned to that same patient. Doing this reliably at a wide-spread scale requires infrastructure, expertise, and precision few can deliver.

ElevateBio was established in 2017 to help biopharma companies industrialize these breakthroughs. Its manufacturing facility in Waltham, Massachusetts operates as a one-stop platform from clinical through commercial manufacturing, capable of supporting thousands of batches annually. Now, ElevateBio is expanding with a new facility in Pittsburgh: a factory of the future leveraging robotics, AI, and digital tools to reduce costs and increase manufacturing scale.

Scaling Cell and Gene Therapy Manufacturing in the United States

In most of the industry, drug discovery and manufacturing operate in silos. Therapies are designed in the lab and only later adapted for manufacturing. ElevateBio eliminates that gap. In addition to manufacturing therapies for biopharma partners, the company has built its own gene editing platform – technology that makes edits directly to the genome with the potential to treat or even cure disease. These tools help partners develop their own therapeutic programs and maximize their chance of reaching patients.

On the manufacturing side, automation, advanced analytics, and process innovation drive down costs while increasing throughput across every batch. This integration matters as these advanced therapies expand into autoimmune diseases, which can have even larger patient populations than oncology indications. The industry needs reliable, U.S.-based manufacturing infrastructure dedicated to advanced therapies that can keep pace – and ElevateBio is building it.

From Waltham to Pittsburgh, ElevateBio is growing its American manufacturing footprint to make production more accessible and globally scalable. Its gene editing services complement the manufacturing platform, giving partners a seamless path from concept through commercialization. The next breakthroughs in genetic medicine won’t come from science alone. They’ll come from the ability to manufacture these therapies reliably, at scale, here in the United States. ElevateBio is building that future, powering the creation of advanced therapies, at a speed the world deserves.

February 26, 2026 by

ElevateBio BaseCamp Pittsburgh

The future of genetic medicine depends on transforming bespoke science into reliable, repeatable manufacturing that can scale globally while controlling costs – ultimately moving advanced therapies from concept to cure. It’s a challenge that echoes Pittsburgh’s own transformation. Just as the city once manufactured the steel that built America, we’re now building the future of healthcare through cell and gene therapy.

It’s this backdrop that shaped our recent discussions with the National Security Commission on Emerging Biotechnology (NSCEB) when they visited BaseCamp Pittsburgh, ElevateBio’s biomanufacturing center at the former industrial site Hazelwood Green, expected to be operational in 2027. During the visit, we discussed NSCEB’s top priorities: scaling and de-risking U.S. biomanufacturing, expanding capacity and workforce talent, and ensuring that American biotechnology innovation remains globally competitive.

ElevateBio is directly aligned with NSCEB’s goals. We leverage deep experience and technical expertise to meet the complex demands of advanced therapy manufacturing, strengthen domestic production capacity, and help fuel the future of medicine.

Industrializing Advanced Therapies

Today, many cell and gene therapies are being produced through unsustainable processes. They hold scientific promise, but manufacturing is expensive and manually intensive. To change this, ElevateBio is industrializing these therapies by implementing standardized, repeatable manufacturing processes and deploying next-generation technologies and automated systems. Taken together, we can enable consistent, large-scale production while simultaneously reducing operational costs. It’s this shift that will move advanced therapies from highly customized, small-batch efforts to sustainable, commercial-ready solutions.

This is what’s required to get these medicines to the patients who need them. By building a proper manufacturing infrastructure, we can expand to treatments for a broad range of conditions, including cancer, autoimmune disorders, and rare genetic diseases.

Expanding U.S. Biomanufacturing Capacity

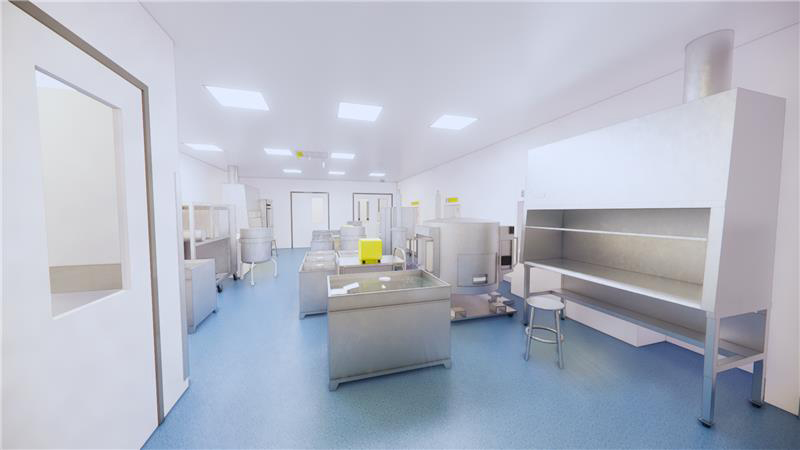
Rendering of a manufacturing suite to be constructed at ElevateBio’s Pittsburgh biomanufacturing facility

Industrializing advanced therapies requires purpose-built facilities capable of supporting programs from preclinical development through commercial scale. BaseCamp Pittsburgh was designed from the ground up to meet these needs, offering flexible manufacturing suites across cell therapy, gene therapy, viral vector, and mRNA platforms.

The facility features a dedicated technology lab designed to evaluate new products and seamlessly integrate innovation into manufacturing operations. Its intentional design aligns materials, workflows, personnel, and information to minimize variability, increase efficiency, and elevate product quality.

Within this lab, we are expanding the traditional role of the manufacturing execution system to accelerate the transfer of new products and innovations into our manufacturing suites. At the same time, it enables more realistic, hands-on training for manufacturing associates – reducing onboarding time while strengthening quality and compliance.

Cultivating the Next Generation of Biomanufacturing Talent

To make this vision a reality, we must invest not only in infrastructure, but also in the people who operate these complex systems. Pittsburgh knows how to build things – and just as importantly, it knows how to train the people who build things. We are building on the strength of Pittsburgh’s leading academic and research institutions – including the University of Pittsburgh and UPMC – and drawing on the region’s deep expertise in AI and automation to advance high-tech biomanufacturing while prioritizing workforce development. Our initiatives include:

  • 170 permanent, full-time positions, many not requiring postgraduate education
  • Partnerships with community colleges to support diverse educational pathways
  • Hands-on, cross-disciplinary training for career changers and early-career talent
  • Collaboration with local industry to identify candidates with existing, transferable skills

These programs reflect ElevateBio’s commitment to building inclusive pathways into advanced biomedical manufacturing, ensuring the next generation of talent can support the growth of U.S. cell and gene therapy production.

Strengthening U.S. Leadership in Biotechnology

Industrializing therapies and building talent sets the stage for reinforcing U.S. leadership in biotechnology. By onshoring production, we can significantly accelerate development timelines, reduce supply chain risk and increase resilience. Leveraging U.S.-based innovation further strengthens the nation’s position at the forefront of global biotechnology. Creating specialized manufacturing hubs like Pittsburgh helps drive the next era of American manufacturing, fostering regional economic growth and technical expertise.

Bringing biomanufacturing to Pittsburgh represents a tangible step in expanding our operations and proving the United States’ ability to produce next-generation medicines reliably and efficiently.

Looking Ahead

The progress at BaseCamp Pittsburgh continues at full pace. Construction has entered its next phase along the Monongahela Riverfront, transforming our long-term vision into physical reality. The facility’s core and shell is complete and meaningful progress has been made on the internal build out. ElevateBio plans to receive a Certificate of Occupancy this year, bringing us closer to making this vision operational.

ElevateBio is proud to extend Pittsburgh’s manufacturing legacy into advanced therapies. By industrializing complex science, strengthening the regional life sciences ecosystem, and training the workforce of tomorrow, we are ensuring the future of medicine is not just invented – but built, accessible, and ready to change lives.

February 6, 2026 by

Throughout my career in cell and gene therapy, I’ve witnessed our industry evolve from scientific possibility to clinical reality. Yet as we scale these transformative therapies, I’m consistently reminded that success hinges not just on the elegance of the science, but on the pragmatic realities of manufacturing.

Choosing a cell therapy contract development and manufacturing organization (CDMO) isn’t a one-time vendor decision. It’s a strategic partnership that will determine whether your therapy reaches patients or joins the sobering percentage of programs that encounter preventable manufacturing setbacks.

Having led process development, manufacturing and global technology transfers early in my career at Bluebird bio, and now serving as Chief Technology Officer at ElevateBio, where I oversee programs across the industry, I’ve identified critical considerations that separate successful partnerships from costly misalignments. The following is a 10-question framework to help inform your decision when choosing a cell therapy CDMO, born from both industry best practices and hard-learned lessons. They’re designed to reveal not what CDMOs promise, but what they can prove and what will determine your program’s success.


1

What are your cell therapy manufacturing success rates?

In my experience, the most revealing metric isn’t what a CDMO highlights in presentations, but their comprehensive performance data. Request their first-time-right manufacturing success rate across all programs. While industry standards hover around 85-90%, exceptional organizations consistently exceed 95%. At ElevateBio BaseCamp, we’ve achieved 98%, though the number itself matters less than the transparency to share it.

Beyond headline metrics, examine deviation rates, failed batches, and out-of-specification results. These indicators reveal the operational consistency that ultimately defines your program. Remember, your batch performance becomes the FDA’s lens into your process control. Inconsistencies documented during early development often resurface as critical observations during BLA review.

2

What experience does your cell therapy manufacturing team have?

Leadership vision matters, but I’ve learned that program success depends on the expertise of those who actually handle your product. In our industry, average GMP manufacturing operator tenure runs one to two years – sufficient for basic proficiency but rarely enough to develop the expertise that distinguishes good from great.

At leading CDMOs, including ElevateBio BaseCamp, you’ll find operators with four to five years or more of specialized experience. Request to meet the manufacturing science, quality, and process development teams who will steward your program daily. Their backgrounds and tenure often predict your program’s trajectory more accurately than executive credentials.

3

Do you offer person-in-plant access during GMP manufacturing?

The question of access reveals much about a CDMO’s operational philosophy. What’s their formal position on person-in-plant presence? Can your team participate in training or observe clean room operations during GMP manufacturing? Is there a limit on the frequency of site visits or are there extensive pre-approvals to do so?

Some organizations restrict access, citing quality or confidentiality concerns. However, I’ve found that transparency typically indicates confidence in both systems and capabilities. At ElevateBio BaseCamp, we actively encourage client collaboration – whether working alongside our technicians during a technology transfer, observing through our in-suite, high-definition cameras, or participating in real-time problem-solving.

4

Can you optimize my cell therapy process or just execute manufacturing?

Nearly half of the programs we’ve worked with at ElevateBio BaseCamp have benefited from process optimization. This isn’t a reflection on our clients’ capabilities, but rather a recognition that cell therapy remains an evolving science where each program presents unique challenges.

Evaluate whether your potential partner maintains dedicated manufacturing science and technology teams that bridge development and production. Request examples of process improvements they’ve implemented. The distinction between a CDMO that merely executes protocols versus one that can scientifically troubleshoot and enhance and industrialize your process often determines whether you’ll navigate challenges successfully or encounter recurring obstacles.

5

Have you passed pre-approval inspection for cell therapy products?

Regulatory readiness extends beyond maintaining compliant systems. It requires demonstrating those systems under the scrutiny of commercial standards. If a CDMO hasn’t yet navigated a pre-approval inspection, investigate what commercial readiness validations they’ve pursued. Third-party certification, like the Initiative for Certification of Manufacturing Capabilities (ICMC™), provide independent verification of quality system maturity.

This consideration carries particular weight given the fact that a significant portion of FDA Complete Response Letters issued between 2020 and 2024 cite manufacturing or quality problems.1 The partnership decisions we make during early development often establish patterns that persist through regulatory review. It’s far more efficient to build commercial-ready rigor from the outset than to retrofit quality systems under regulatory pressure.

6

What’s your standard technology transfer timeline for cell therapy programs?

Technology transfer represents one of the most underestimated risks in our industry. I’ve seen programs lose momentum – and sometimes commercial competitiveness – due to protracted or failed transfers. Ask potential partners about their recent track record: How many transfers have you completed successfully over the past three years? What percentage met original timelines versus requiring extensions?

The financial and reputational costs of a failed CDMO relationship extend well beyond direct expenses. Programs can lose years and deplete resources that can’t be recovered, leaving teams to navigate compressed timelines with diminished funding. Historical performance, particularly with programs similar to yours, offers the clearest indicator of future success.

7

Can you scale cell therapy manufacturing from Phase 1 to commercial?

Success in cell therapy can paradoxically create its own challenges if your manufacturing partner lacks scaling capability. I’ve observed promising programs stall not from clinical failures but from inability to demonstrate manufacturing consistency at increased scale, a regulatory requirement that catches many teams unprepared.

Request concrete evidence of scaling experience: documented capacity expansion plans, not aspirations. Understand whether capacity is reserved for existing partners or subject to competitive allocation when demand peaks. Most importantly, verify they’ve successfully transitioned programs from clinical-scale production to commercial volumes while maintaining the consistency regulators require. Your manufacturing partner’s growth trajectory must align with your program’s ambitions.

8

What regulatory expertise and infrastructure do you provide for BLA submissions?

The FDA doesn’t just review your final product – they review your entire journey and product lifecycle. Can your CDMO demonstrate successful navigation of FDA feedback? How many INDs and BLAs have they actually supported? Do they have former FDA staff who understand how reviews really work, not just theoretical knowledge?

Equally critical is the digital infrastructure supporting your regulatory submissions. What systems ensure the data integrity FDA demands? Electronic batch records, integrated quality management systems, and comprehensive audit trails are regulatory requirements. Review the systems your CDMO has in place and ask for specific examples of how they’ve managed inspection observations to turn potential issues into approvals. The difference between a CDMO that reactively responds to regulatory requirements and one that proactively anticipates and addresses them often determines whether your program proceeds smoothly or encounters unexpected delays.

9

Was your facility purpose-built for cell therapy, and how does your team integrate new technologies?

There’s a fundamental difference between facilities designed for cell therapy and those retrofitted from other modalities. ElevateBio BaseCamp was built with FDA input specifically for multimodal, multiproduct production of cell, gene and mRNA therapies, with infrastructure optimized from material flow to contamination control and environmental monitoring. In contrast, so-called “flexible” facilities originally designed for stable molecules or well-characterized biologics are often compromised across these requirements.

Equally important is how that infrastructure evolves. The cell and gene therapy field evolves rapidly, yet many CDMOs hesitate to integrate innovations that could benefit their clients’ programs. Ask for specific examples of recently implemented technologies. How do they evaluate new automation or analytical methods? Do they have a technology development lab where innovations can be tested without risking GMP production? At ElevateBio BaseCamp, we’ve implemented more than ten new technologies in the past year alone, from automated processing platforms to advanced analytical methods. The willingness and capability to evolve with the science often distinguishes partners who will advance your program from those who might constrain it.

10

What are your sustainability commitments and environmental certifications?

Many biopharmaceutical companies look for environmental commitments from their suppliers, becoming just as important as quality systems in vendor selections. Ask how your CDMO considers environment and occupational health certifications. At ElevateBio BaseCamp, we pursued International Organization for Standardization (ISO) 45001 and 14001 certifications early, recognizing that our commercial partners would eventually require this level of rigor from their supply chain.

As we scale cell therapies toward broader patient populations, demonstrating sustainable manufacturing practices becomes part of our collective responsibility to deliver these treatments responsibly.


These questions are designed to reveal which partners truly understand the complexity of cell therapy manufacturing. The right CDMO won’t hesitate to share specific metrics, provide references, or open their doors for inspection. They’ll welcome these questions because they’ve already built their operations around answering them.

At ElevateBio BaseCamp, we built our operations specifically to address these challenges. From our purpose-built facilities to our experienced team and commercial scale, we welcome these tough questions.

Learn more about ElevateBio BaseCamp’s approach

References: 

  1. Slabodkin, Greg. “FDA’s CRLs Reveal 74% of Applications Rejected for Quality, Manufacturing Issues.” Pharma Manufacturing, 14 July 2025, www.pharmamanufacturing.com/all-articles/article/55302937/fdas-crls-reveal-74-of-applications-rejected-for-quality-manufacturing-issues.

Mike Paglia, Chief Technology Officer

Michael Paglia is the Chief Technology Officer at ElevateBio, a technology-driven company commercializing its enabling technologies, manufacturing capabilities, and industry-leading expertise to accelerate the development of genetic medicines to treat human diseases. He has more than two decades of experience in facility design, start up, and operations ensuring the highest standards of quality, safety, and regulatory compliance. 

At ElevateBio, Michael led the design, construction and operations of the BaseCamp manufacturing facility that was recognized as the Facility of the Year, Operational Excellence by International Society for Pharmaceutical Engineering (ISPE) in 2021.  Michael established the process development and manufacturing capability and leads manufacturing operations, CMC regulatory, process/analytical development, and the advancement of innovative process technologies. 

Prior to ElevateBio, Michael was the Vice President of CMC Operations at Oncorus responsible for the development and manufacturing of novel genetically modified oncolytic herpes virus for the treatment of cancer and prior to that, Head of Technical Operations, Cellular Process Development and Manufacturing Operations at bluebird bio where he led the early process development, manufacturing, and global technology transfer of four approved genetically modified autologous cell therapies.  Early in his career at Tolerx, Michal lead process development, and late-stage manufacturing of novel therapeutic antibody products designed to treat patients by reprogramming the immune system.

Michael received his undergraduate degree from Providence College and a Master’s of Science in Biochemistry and Cellular Biology from the University of New Hampshire where he was honored with the Distinguished Alumni Award from the College of Life Science and Agriculture (COLSA) in 2023 for his career guidance and ongoing initiatives in COLSA to enhance STEM workforce development initiatives.
February 6, 2026 by

In the race to bring transformative cell and gene therapies to patients, speed often dominates early decision-making but industry data reveals a significant trend: between 2020 and 2024 a significant portion of FDA Complete Response Letters (CRLs) issued by the U.S. Food and Drug Administration involve manufacturing and quality issues.1 This industry trend is also reflected specifically in cell and gene therapy, where complex processes and novel modalities amplify the risk. These setbacks are rarely caused by last-minute missteps. More often, they trace back to decisions made years earlier during preclinical and Phase 1 testing when programs are under pressure to move fast and reduce costs.

A Predictable Pattern of Late-Stage Setbacks

Across the industry, the same challenges continue to emerge late in development. These findings don’t arise overnight – they expose gaps that were embedded in development programs years earlier.

The consequences surface at the worst possible moment: when a company is advancing toward approval, investor expectations are highest, and five or more years of development – and significant capital – have already been invested. The results are major approval delays, immense unplanned costs, and challenges that can fundamentally alter a company’s trajectory.

Cell Therapy Intensifies the Challenge

While these statistics highlight industry-wide trends, cell therapy adds unique challenges that magnify these risks. In this space, early decisions carry disproportionate weight: deficiencies in process design or scale-up can ripple through development and delay approval, even years later.

These risks tend to play out in consistent ways across programs, pointing to key areas that must be managed carefully to ensure successful development.

Critical issues to avoid:

Unresolved CMC and facility readiness issues, with critical details missing from Chemistry, Manufacturing, and Controls (CMC) packages, and manufacturing sites not fully prepared for FDA inspection

Assays not built for late-stage demand, often revealing limitations because they were designed for early research rather than commercial scale, robustness, and regulatory expectations

Product quality and manufacturing success-rate challenges, where teams struggle to consistently produce product that meets specifications, particularly around viability, stability, and other critical quality attributes

Difficulty scaling manufacturing, where processes that work at early stage can fail under commercial demand, making it hard to demonstrate comparability, reproducibility, or consistent performance

Building Success from the Start: ElevateBio Addresses the Root Causes

At ElevateBio, we’re focused on advancing the field of cell and gene therapy by combining genetic medicine technologies with manufacturing scale and expertise. So, we understand that manufacturing cell therapy is inherently complex, requiring robust processes, careful planning, and rigorous quality systems from the very start. This requires the right processes, the right people, and a quality-first mindset embedded from day one, so we can help our partners avoid the costly mistakes that set their programs back.

ElevateBio BaseCamp® is dedicated to the development and manufacturing of genetic medicines to address these challenges. Designed to be an integrated part of our partners’ development and approval journey, BaseCamp provides the foundation needed to withstand late-stage scrutiny and accelerate time to patients.

What sets ElevateBio BaseCamp apart:

  • A world-class team with proven experience manufacturing and releasing complex cell and gene therapy products
  • Expanding commercial manufacturing infrastructure engineered for reliability, scale, and regulatory readiness
  • Deep product understanding, supported by regulatory expertise and advanced analytical capabilities
  • A culture of quality and collaboration that prioritizes speed with accuracy, transparency, and true partnership

This combination matters because ElevateBio has already solved the problems others are discovering too late. Our partners benefit from established systems, extensive experience, and an operational model designed to anticipate regulatory and manufacturing challenges.

Manufacturing Setbacks are Not Inevitable

Many issues stem from rushing early development, choosing the wrong partners, or re-learning lessons the industry already knows.

The promise of genetic medicines is real. These therapies are transforming care for diseases once considered untreatable. But realizing that promise requires treating manufacturing as a strategic driver, not a downstream function. In cell and gene therapy, regulatory success is shaped years before submission and depends on partners with the right processes and quality systems in place from the start. That is what ElevateBio provides: the experience, infrastructure, and commitment to quality needed to turn scientific breakthroughs into approved therapies – and ultimately deliver them to patients who are waiting.

Learn more about ElevateBio BaseCamp’s approach

References: 

  1. Slabodkin, Greg. “FDA’s CRLs Reveal 74% of Applications Rejected for Quality, Manufacturing Issues.” Pharma Manufacturing, 14 July 2025, www.pharmamanufacturing.com/all-articles/article/55302937/fdas-crls-reveal-74-of-applications-rejected-for-quality-manufacturing-issues.

Cindy Riggins, Ph.D., Vice President, CMC Regulatory Affairs

Cindy Riggins, Ph.D. is Vice President, CMC Regulatory Affairs at ElevateBio. Cindy started her career in cell and gene therapies in 2001 at FDA/CBER as a post-doctoral fellow studying xenotransplantation and later transitioning to product reviewer for various cell therapy products. After leaving FDA in 2008, she has been involved in development of monoclonal, cell, and gene therapies through CMC Regulatory Affairs roles at AstraZeneca, Novartis, Autolus and ElevateBio. She was part of the regulatory team at Novartis responsible for submission and approval of Kymriah®, the first gene therapy product approved in the USA.

November 26, 2025 by

Over the past decade, our industry has witnessed the scientific promise of cell and gene therapies. Patients with rare diseases or hard-to-treat diagnoses now have new treatment options harnessing human cells and genes to alter disease. But the accessibility of these therapies the industry has developed remains constrained not by what’s biologically possible, but how they are designed and manufactured.

The field has reached an inflection point. We’ve demonstrated the scientific foundation and its curative potential. But to make advanced therapies sustainable as a pillar of medicine, we must make them more accessible. The companies that will define cell and gene therapy’s future will be those who can eliminate the distance between top science and efficient manufacturing.

Integration of Manufacturing and Therapeutic Design

Traditional small molecule drug development has very siloed development pathways: a therapeutic is designed and developed by one team and then manufactured by another. This approach is challenging in cell and gene therapy, often leading to delays, setbacks, or even outright failures. We built ElevateBio to solve this problem with a new approach, one in which therapeutic design and manufacturing operate as an integrated ecosystem.  

ElevateBio BaseCamp, our cGMP manufacturing business, goes beyond a traditional CDMO. We bring together expertise, advanced technologies, and state-of-the-art facilities to serve as a skilled partner to biopharmaceutical companies. This includes in-house manufacturing, process and analytical development, and quality control teams, all working in parallel to achieve tighter coordination and faster turnaround times. BaseCamp has industrialized genetic medicine manufacturing, achieving a 98% batch success rate across advanced programs.

Yet sustaining this success – and expanding it across new modalities – requires more than technical excellence alone.

Designing for Manufacturability from Day One

The future of cell and gene therapy depends on therapies designed with manufacturability in mind from the start – and into every stage of design. That’s why our team of process development scientists are embedded in design conversations early, creating commercial-ready processes in parallel with therapeutic development. Manufacturing insights flow back to inform molecular engineering in real time.

This includes integrating compact constructs and delivery systems engineered for both efficacy and efficiency. We apply scale-down and scale-up models to optimize performance, ensuring processes are fully scalable to GMP manufacturing and capable of meeting global demand.

We take the same approach through ElevateBio Life Edit, our gene editing technologies and R&D business. When our teams develop gene editors across all modalities, manufacturability is a design criterion from day one – not a constraint discovered in late-stage clinical trials. And by having BaseCamp and Life Edit sit alongside one another, we’re ensuring the latest manufacturing developments and insights flow back to inform R&D – and vice versa.

A Foundation for an Industry to Prosper

Looking beyond the science, what does a sustainable cell and gene therapy ecosystem require?

It’s more than better therapeutics. We need more treatment centers, expanding from dozens to hundreds for better patient access. The industry needs new commercial models that make advanced therapies economically viable for health systems. We need a whole new infrastructure where cell and gene therapy can become the standard of care for previously untreatable conditions.

But that infrastructure can’t be built upon unreliable manufacturing. We as an industry need to build a strong foundation – one built by designing, optimizing, and validating processes that reliably move therapies from bench to bedside. Without that foundation, the ecosystem simply can’t scale. And the window to build it is narrowing.

CAR-T is expanding into autoimmune indications with patient populations 10 times larger than oncology. In vivo therapies are advancing as new-generation modalities are adding layers of complexity. To support this growth, the field needs manufacturing designed for reliability and scale from day one.

Building What Comes Next

The field now needs the operational discipline and integrated thinking to deliver on that promise at population scale.

The therapies we’re developing today have the potential to transform millions of lives. But only if we build the systems to make them accessible, reliable, and sustainable.

At ElevateBio, we’re building that foundation by combining BaseCamp’s manufacturing platform with Life Edit’s R&D capabilities – and embedding therapeutic design expertise throughout. By doing so, we’ve created an integrated approach that’s building cell and gene therapy’s future and making a tangible impact for patients worldwide.

May 23, 2025 by

Drug Target Review recently published a byline article featuring LETI-101, Life Edit’s development candidate for Huntington’s disease (HD). HD is a rare, inherited neurodegenerative disorder affecting approximately 41,000 people in the United States, with another 200,000 at risk. LETI-101 uses Life Edit’s CRISPR technology for allele-selective editing, offering a novel approach to potentially treating this devastating condition. What sets LETI-101 apart is its precision targeting strategy. Rather than directly targeting the disease-causing CAG repeat expansion, LETI-101 targets single nucleotide polymorphisms (SNPs) that allow us to distinguish between mutant and healthy copies of the gene.

“A transformative aspect of our LETI-101 approach is that it’s designed as a one-time treatment that could provide long-lasting benefit without the need for repeated administration,” explains Dr. Amy Pooler, SVP of Research & Development at Life Edit. “Unlike other therapeutic modalities being developed for HD that would require ongoing treatment to maintain efficacy, our gene editing therapy is intended to make a permanent, precise modification to the DNA itself.”

Read the full article “Allele-selective gene editing: a breakthrough in Huntington’s disease treatment” on Drug Target Review.

April 4, 2025 by

In March 2022, Beam Therapeutics and ElevateBio embarked on a groundbreaking partnership to manufacture BEAM-101, an investigational base editing therapy for sickle cell disease. The therapy represents a novel approach to treating this devastating condition by mimicking genetic variants found in individuals with hereditary persistence of fetal hemoglobin. To advance this potentially transformative treatment to clinical trials, Beam turned to ElevateBio’s expertise in cell and gene therapy production, combining rigorous quality standards with the ability to move at unprecedented speed.

The Challenge

Manufacturing gene-edited cell therapies presents unique complexities beyond traditional cell therapy production. Base editing, like all gene editing cell-therapy technologies, requires careful process control to maintain editing efficiency and cellular viability. Traditional manufacturing approaches, optimized for other cell therapies, needed significant adaptation for this novel technology. Additionally, Beam faced urgent timeline pressures – they had identified potential clinical trial participants and needed to accelerate their manufacturing readiness without compromising quality.

“Base editing represents a new frontier in genetic medicine, and its manufacturing demands match that innovation,” says Brian Riley, chief manufacturing officer of Beam. “We needed a partner who could not only handle the technical complexity but move at the speed required to help us reach waiting patients.”

Pioneering Base Editing Technology

Beam’s proprietary base editing technology allows for precise genetic modifications without making double-stranded breaks in the DNA. BEAM-101 leverages this precision to mimic genetic variants seen in individuals with hereditary persistence of fetal hemoglobin – a natural condition that protects against the effects of sickle cell disease. This approach represents a potentially transformative treatment option, offering hope for a one-time therapy that could provide lasting benefit to patients.

Elevate Bio

BaseCamp’s End-to-End Manufacturing Capabilities

ElevateBio BaseCamp is a state-of-the-art process development and cGMP manufacturing facility designed to accelerate the development of cell and gene therapies. BaseCamp provides comprehensive manufacturing solutions, combining advanced technical capabilities with experienced program management to support partners from early development through clinical trials. The facility’s integrated approach to process development and manufacturing enables rapid technology transfer and scaling of complex genetic medicines. This comprehensive platform enables rapid process optimization and seamless technology transfer, crucial capabilities for novel therapeutic modalities like base editing.

The Approach: Innovation at Speed

Beam partnered with ElevateBio’s BaseCamp to define a plan for clinical supply, which ultimately led to an accelerated tech transfer to support BEAM-101’s path to clinical trials.

BaseCamp’s comprehensive strategy combined technical expertise with dedicated program management. The seamless integration of program management and the Manufacturing, Science and Technology (MSAT) group provided end-to-end oversight for technology transfer activities, while parallel manufacturing readiness and analytical method transfer activities compressed traditional timelines. The team leveraged a risk-based approach along with input from the process development group for successful translation of the novel base editing process into GMP manufacturing. This was supported by an accelerated training and qualification program that maintained the highest quality standards.

“We recognized that standard technology transfer approaches would not meet the timeline requirements,” explains Mike Paglia, Chief Technology Officer, ElevateBio BaseCamp. “Our team developed an integrated strategy leveraging our development, manufacturing and quality teams that maintained the highest level of quality while dramatically accelerating the timeline of the technology transfer of the Beam-101 process to ElevateBio BaseCamp.”

Michael Paglia,
Chief Technology Officer, ElevateBio BaseCamp

The results were immediate. Within four weeks, the teams completed their first training run. A second run followed within a month, and engineering runs began shortly after. This unprecedented speed came without sacrificing thoroughness – the teams transferred nine critical assays, completed comprehensive qualification, and maintained rigorous quality standards throughout.

Technology Innovation

The partnership demonstrated the value of close collaboration between therapy developers and manufacturing partners in establishing robust production processes. The team’s experience in transferring and scaling complex cell therapy processes helped create effective workflows and quality control approaches that supported the successful manufacture of BEAM-101.

Results: Breaking New Ground

The partnership achieved several breakthrough milestones in manufacturing excellence. Most notably, the team completed technology transfer two months ahead of schedule – while maintaining exceptional quality standards. The manufacturing process demonstrated consistent editing efficiency and reliable production parameters, establishing a repeatable model for future programs.

Key achievements include:

  1. Completion of comprehensive tech transfer ahead of schedule.
  2. Achieving successful site-to-site comparability to Beam’s in-house manufacturing facility
  3. Proudly supplying patient doses for Beam’s BEACON Phase 1/2 clinical trial evaluating BEAM-101 in adult patients with sickle cell disease

Beyond these immediate results, the collaboration delivered strategic value that extends well beyond BEAM-101. By working with ElevateBio, Beam successfully demonstrated the manufacturing scale-up of its base editing technology – a critical milestone in proving the commercial viability of any novel therapeutic platform. This demonstration provides a blueprint for future development programs across Beam’s pipeline, potentially accelerating the timeline for bringing additional base editing treatments to patients.

“The speed and quality of this collaboration exceeded our expectations,” says Giuseppe Ciaramella, Ph.D., president of Beam. “BaseCamp didn’t just manufacture our therapy – they helped us establish a blueprint for base editing manufacturing.”

Giuseppe Ciaramella, Ph.D.
President, Beam Therapeutics

The successful manufacturing of BEAM-101 represents more than one company’s achievement – it demonstrates the feasibility of rapidly scaling novel genetic medicines. As the field of genetic medicine advances, manufacturing remains a critical determinant of success. The manufacturing platform and processes developed through this collaboration create a pathway for future base editing therapies, potentially accelerating the development of treatments for other genetic diseases.

February 14, 2025 by

Cell therapies have emerged as a transformative tool of modern medicine, offering unprecedented potential to treat and cure a wide range of diseases. Engineered cell and gene therapies are able to address the etiologic genetic mutation or eradication of the disease-relevant cellular compartment, with profound improvements in clinical outcomes. From immune-based approaches like T cell therapy for cancer to regenerative applications utilizing stem cells, cell therapies are redefining the boundaries of treatment modalities. 

Gene delivery technologies enable the introduction, deletion, or modification of genetic material within cells, equipping them with novel therapeutic properties or optimizing their natural capabilities. From viral vectors such as lentiviruses and adenoviruses to non-viral methods like electroporation and lipid nanoparticles, these technologies form the backbone of genetic engineering in cell-based treatments. 

The quality control measures that underpin the development and commercialization of these promising therapeutics are key to furthering them within the larger biopharmaceutical pipeline. In particular, potency assays are central to this pursuit, as these analytics are crucial to ensuring product consistency, efficacy, and safety. The potency of a product is the specific ability or capacity of a product to achieve a defined biological effect. Potency assays are quantitative measures of biological activity and are typically assessed in vitro. 

In the case of CAR or TCR T cell products, both the vector to deliver the gene of interest (GOI) and the gene-modified T cell drug product require potency assays to be in place to support product release and stability. By adopting a phase-appropriate yet prospectively considered approach to potency development as early as possible in a process, organizations can arrive at a potency control strategy that improves the foundational understanding of a product’s quality and consistency and results in a strategy that will be suitable for a marketing application while not jeopardizing the use of valuable clinical data needed to support the safety and efficacy assessment for the application.

Genetic medicine potency assays

The development of potency assays can be challenging due to the complex nature of cell and gene therapies and the lack of standardized methods in the broader development space. Development of suitable and robust potency methods requires plenty of development data and correlation from orthogonal readouts. During early phases of drug development, a potency assay can be a quick and simple method suitable for the phase. However, through the course of drug development, potency assays often require several rounds of iteration and maturation, including implementation of controls and standards. Moreover, the functional potency assays that support a marketing application’s overarching potency strategy must be able to effectively measure a product’s mechanism of action (MOA) or biological function. For many complex products, the understanding of the drug MOA evolves through the course of development. It is therefore recommended that the potency work should start early during development.

The assay development can come at a significant cost as the assays may require several custom reagents, including the need for establishing cell banks and reference materials. Developing a potency strategy for genetic medicines is often challenging for the companies pioneering these treatments, many of whom are working with small teams, constrained resources, and competing priorities throughout development. 

Regulatory expectations for potency assays

The existing successes of CAR and TCR T cell products mean that the regulatory expectations around these products are reasonably well documented1,2,3 — including that potency assays should:

  • Reflect biological effects that represent the proposed clinical MOA 
  • Characterize a product well enough to identify and evaluate the impact of process changes
  • Enable operators to establish criteria for stability and comparability during process changes, improvements, and lot release. 

Pre-Clinical Development to FIH

  • Establish proof of concept
  • Initiate development of multiple readouts: genetic and protein
  • Semi-quantitative with phase-appropriate specificity and sensitivity
  • Evaluate suitability for in vitro and animal model testing

Later Phases of Clinical Development To Pivotal

  • Refine assays for quantitative readouts based on early clinical data: Identify Reference standards and critical reagents
  • Develop MoA functional potency
  • Qualify assays for accuracy, precision, and robustness
  • Assess suitability for later phases: Establish acceptance criteria

Toward Commercial Filing

  • Further optimize assay based on expanded clinical data
  • Validate with larger sample size and routine handling conditions
  • Finalize documentation for regulatory submission
  • Confirm method acceptance criteria

Figure 1: An overview of the key considerations for a phase-appropriate potency strategy. 

The regulatory agencies suggest potency assays be in place even during the initial phases of development so that, by the time the product has moved into pivotal efficacy studies, quantitative potency assays that measure MOA-reflective biological activity are required for lot release and stability. The latest FDA guidance1 emphasizes a lifecycle approach to potency that is grounded in quality risk management, where potency tests are considered throughout the product lifecycle, from product development all the way to product licensure, and can adapt with gained knowledge of mechanism of action and assay experience.

Although these requirements are widely acknowledged, many companies run into snags early when it comes to approaching potency. For example, an organization can focus exclusively on potency for the drug product without recognizing that the vector is also considered a critical component that furnishes a pharmacological activity to the drug product and should include testing of biological activity. Or an organization may not have the bioassay development expertise or the regulatory experience to develop the potency control strategies in a phase-appropriate manner. Engaging with a full-service CDMO with sophisticated analytical capabilities, expertise, and infrastructure can help expedite the development of potency control strategies.

Furthermore, while early development potency lot release assays can be less stringent “litmus tests,” these analytics are likely to eventually need to have two-sided acceptance criteria. Developers must also consider the type of statistical analysis they perform, such as parallel line analysis for more complex assays at later phases to demonstrate similarity to a reference material. Establishing and maintaining a reference material is ideal as these provide a consistent point of reference to compare the biological activity of a drug product or substance, ensuring accurate and reliable relative potency measurements throughout development. 

Companies that deprioritize development of methods to measure biological potency until later phases of development risk falling behind in maturing their assays effectively and can encounter regulatory and technical setbacks as a program progresses. This can make it hard for organizations to pinpoint challenges, even for those that have retained earlier samples for testing, as the quality and stability of these retains cannot be assured. 

Creating a balanced potency assurance strategy 

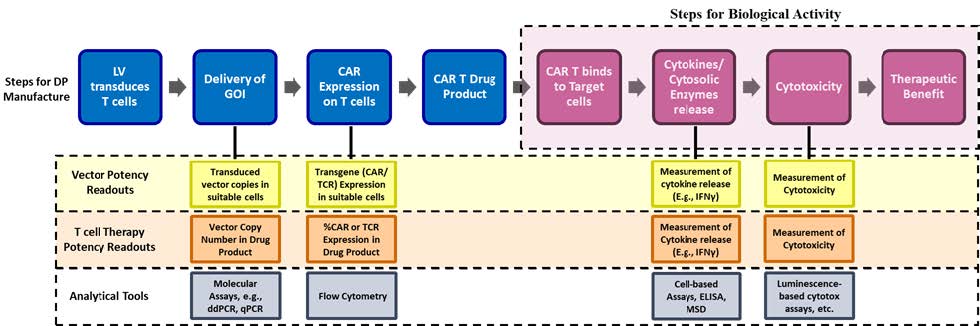
CAR and TCR T cell therapies affect target cells in an antigen-specific manner using multiple mechanisms, and therefore the use of orthogonal methods is recommended. A CAR or TCR T cell product is made by delivering the GOI using a suitable vector, e.g., a lentiviral vector (LVV). Upon GOI delivery, the engineered receptor is expressed on the T cells that can bind to specific antigens on target cells (e.g., cancerous cells). When the therapeutic cells interact with target cells via the engineered CAR or TCR, intracellular signaling cascades within the DP cells leads to the release of pro-inflammatory cytokines, cytolysis of the target cells, and expansion and proliferation of the engineered cells. These are key indicators of T cell activation. Thus, interferon gamma (IFNγ), a pro-inflammatory cytokine, serves as a critical downstream marker in this cascade, making it a relevant attribute for the MOA and quantifiable readout of CAR function4. 

When measuring potency for these complex therapeutics, a set of potency assays has been well validated for these applications (Figure 2). They include:

  1. Measuring the delivery and integration of GOI at genetic level: This can be done by using molecular techniques like ddPCR or qPCR.
  2. Measuring the expression of transduced GOI at a protein level: Transgene expression can be measured using flow cytometry to quantify the percentage of cells expressing the CAR.
  3. The biological activity of the GOI can be measured by quantifying cytokine release using cell-based assays such as ELISA, ELLA, MSD, or flow cytometry. 
  4. The biological activity of the GOI can also be measured by quantifying the killing of target cells using cell-based assays leveraging luminescence or flow cytometry.

Figure 2: A simplified overview of the gene-edited cell therapy manufacturing process and potency assay strategy based on key process steps.

In the example of a CAR or TCR T cell therapy, expression assays for the GOI are necessary for understanding potency for early-stage processes. Expression assays alone do not offer insight into the biological function of the cell product, however. This is where assays like those used for cytokine release or cytotoxicity are integral to a program and why at least one should be incorporated early, even if they are not used as qualified release assays at this nascent stage of development. These tests require more up-front work for method development — establishing acceptance criteria and creating controls and reference standards — but they can offer greater insight that is indispensable in the long term. 

Ideally, biological potency assays can be introduced early in development to gain product and assay knowledge critical to enabling the most appropriate methods and acceptance criteria for release and stability testing during pivotal clinical trials. Additionally, these assays are extremely useful to have in analytical comparability studies, including for changes introduced in early development. 

The vector potency determination also takes a similar approach of analytical readouts (Figure 2). During the initial phases of development for these therapies, the primary focus is on verifying the ability of the vector to successfully deliver the GOI into representative cells. This approach typically employs transduction of a target cell line by the vector. The transduced cells are cultured, harvested, followed by PCR amplification of the integrated provirus sequence, offering a precise measurement of the delivered gene copy. The functional vector potency readout is designed to demonstrate the ability of the vector to generate a biologically active CAR or TCR T cell based on readouts like cytokine release.

Key considerations during development of potency methods


Method development and optimization for these therapies require a systematic approach to ensure robust and reproducible processes that deliver high-quality therapeutic products. The molecular methods measuring drug product potency are based on accurate and precise quantification of cells with integrated vector based on a PCR-based readout. The development and qualification of a molecular assay are relatively straightforward and focus mainly on optimizing the primers, probe and PCR conditions, plus ensuring appropriate method controls. Similarly, the measurement of %CAR or %TCR -positive cells in DP using flow cytometry requires identification of appropriate antibodies and optimizing the staining conditions and gating strategy.

The biological potency for T cell product can be based on in vitro cell-based methods aimed at measuring cytokine release and/ or cell cytotoxicity. The method setup requires activation of T cell drug product by co-culture with antigen-presenting target cells or incubating with target antigen. Establishing the critical reagents, most importantly the target cell line or target antigen, is the first step to developing a robust method. Several pros and cons must be considered when finalizing the choice for the activation step as it must demonstrate consistent response in the potency readout. The target cell lines must be comprehensively tested to confirm cell viability, genetic stability, and expression of the specific antigen at appropriate levels. 

Optimization of the method variables such as cell seeding density, effector-to-target cell ratio during co-culture, duration of culture, etc., are equally critical to building a robust method. Part of the puzzle also requires identifying and optimizing a suitable analytical readout. For a cytokine release potency, the typical analytical tools include ELISA, ELLA, MSD, etc. The qualities influencing the choice of readout often include accuracy, precision, and robustness of the method, operator hands-on time, degree of automation, availability, and cost of suitable kits and reagents. 

The analytical tools and readouts used for vector potency measurement are similar to the T cell therapy product, but the key difference is the design, setup, and reportable results determining potency. A vector potency setup typically uses representative non-transduced cells, either derived from healthy donors or a suitable cell line, that are transduced in small-scale format by titrating vector test article. Upon completion of culture, the cells are harvested and tested to measure the transduction ability of the vector, plus downstream function, such as cytokine release, of the delivered GOI. 

Generating and characterizing cell banks is critical as it serves as a consistent and reliable starting material for transduction. Equally critical is bridging and demonstrating comparability of cell banks when needed for ensuring method consistency through the product’s lifecycle. To assess vector dose response, dose titration studies are conducted to correlate vector concentration with functional outcomes, such as transgene expression or biological activity4. Variables such as multiplicity of infection (MOI) range, transduction conditions such as exposure time, and media composition, are carefully evaluated to optimize gene transfer and maintain cell health. Post-transduction, harvest conditions, including timing and cell viability, must be standardized for a robust and consistent method performance. 

Summary

Overall, early development of robust potency strategies is crucial for ensuring the clinical and regulatory success of advanced therapies. By integrating phase-appropriate assays and continuously refining methods based on evolving understanding of mechanisms of action, developers can mitigate risks and improve product consistency. The establishment of reliable reference materials and the use of orthogonal testing approaches likewise provide critical insights into a product’s biological activity, facilitating smoother regulatory submissions. Ultimately, a well-defined potency strategy supports the timely delivery of quality, safe, and effective therapies while minimizing setbacks and aligning with regulatory expectations. 

Many sponsors face challenges developing a potency control strategy specifically for cell and gene therapies due to the complexities in understanding and measuring the biological effect of the products. Additionally, smaller, younger companies may not have the required resources and expertise, or a larger organization may be working on more traditional modalities and not have the CGT experience. With extensive experience across a range of modalities—including genetically modified cell therapies, gene editing, and viral and non-viral vectors—ElevateBio ensures that potency assays evolve appropriately throughout the development lifecycle, from preclinical stages to first-in-human trials and beyond, supporting critical milestones toward commercial filing. This phase-appropriate, data-driven approach enables companies to meet regulatory requirements while optimizing the consistency, safety, and efficacy of their therapies.

References: 

  1. U.S. Food and Drug Administration. (2023). Potency Assurance for Cellular and Gene Therapy Products: Draft Guidance for Industry. Retrieved from https://www.fda.gov/regulatory-information/search-fda-guidance-documents/potency-assurance-cellular-and-gene-therapy-products
  2. European Medicines Agency. (2023). Guideline on quality, non-clinical and clinical aspects of medicinal products containing genetically modified cells (Revision 1). Retrieved from https://www.ema.europa.eu/en/documents/scientific-guideline/guideline-quality-non-clinical-and-clinical-aspects-medicinal-products-containing-genetically-modified-cells-revision-1_en.pdf
  3. U.S. Food and Drug Administration. (2011). Guidance for Industry: Potency Tests for Cellular and Gene Therapy Products. Retrieved from https://www.fda.gov/files/vaccines,%20blood%20%26%20biologics/published/Final-Guidance-for-Industry–Potency-Tests-for-Cellular-and-Gene-Therapy-Products.pdf.
  4. Kiesgen S, Messinger JC, Chintala NK, Tano Z, Adusumilli PS. Comparative analysis of assays to measure CAR T-cell-mediated cytotoxicity. Nat Protoc. 2021 Mar;16(3):1331-1342. doi: 10.1038/s41596-020-00467-0. Epub 2021 Feb 15. PMID: 33589826; PMCID: PMC8064272.

September 27, 2024 by

When ElevateBio was founded, we knew that the cell and gene therapy (CGT) landscape would need significant technological innovation to realize the true potential of these complex therapeutics – not only to design them and better understand what makes them safe and effective, but also to identify more robust and cost-effective ways to manufacture them at scale. Some of the required technologies were already available, but further improvements and developments were needed to enable process automation and aseptic processing. This meant we needed to think outside of the box to build the technological future of CGTs and truly bring about the sea change we envision for the field to change the future of medicine.

ElevateBio can think outside the box because we don’t focus on a single therapeutic modality and its associated technology. Rather, we span the entire spectrum of genetic medicine modalities and have visibility and access to a wide range of tools and technologies deployed across the biotech sector, as well as other sectors. We also have visibility to many of the common challenges and issues facing CGT development and manufacturing, which provides us with insights on the types of technological innovations that may be needed most. This broader perspective informs how we evaluate tools and technologies and apply them to the needs of CGTs.

To support our continued technology development efforts, we created our Emerging Technology Lab for cell and gene therapies. This lab was carved out with a focus on new technologies to yield high-quality, potent therapies, using more robust, cost-effective and scalable manufacturing approaches, so that more patients can benefit from genetic medicines.

What we do

Technological innovation is critical to the development and manufacturing of CGTs because they are far more complex than small molecules or biologics – and their complexity is only increasing as the development landscape evolves.

To stay ahead of these increasing product complexities, our emerging technology lab is a beta testing center for new devices, reagents, analytical technology, sensor technology, automated and digital control systems, with a focus on addressing common cell and gene therapy manufacturing challenges.

As a technology-driven company, we are constantly evaluating new tools on behalf of vendors, partners, and industry collaborators. Many of the technologies we evaluate were designed and built for just one, very specific purpose that applies to a single therapy or approach. But from our 30,000-ft viewpoint of the entire genetic medicine sector, we can look at existing tools and recognize how they could be used in new and different ways, such as redeploying the robotics used in factory warehouses for process automation. We present the technology to the whole team – with its breadth of experience – and ask: How could we use this piece of technology beyond what it was designed it to do?

Table 1 highlights several examples of our evaluations in terms of the novel or emerging technology type, the process unit where it would be used, and the advantages or improvements it could offer.

What makes our approach unique

ElevateBio is certainly not alone in evaluating emerging technologies for genetic medicines.

However, companies tend to focus on technology development according to what is relevant to the segment(s) of the product life cycle they occupy: a company developing therapies for first-in-human studies is probably not developing a commercial-ready automated and closed manufacturing process; CDMOs and other companies that specialize in GMP manufacturing run the risk of dedicating most of their time to establishing consistent and efficient operations and not investing enough time and effort in evaluating novel technologies for improving product design and process development. By contrast, ElevateBio occupies the entire product life cycle, and our expertise is unique because of the previous experience some of our team members have in developing some of the early CGTs that are now commercially approved, as well as our ongoing exposure to many different products and therapeutic modalities at all phases of development.

Therefore, ElevateBio’s focus is more technology-intensive than other companies because we occupy the entire continuum of genetic medicines development. Our end-to-end technological capabilities and expertise allow us to pursue technology development in ways that are unmatched by other companies in the sector. Our technology development efforts also have a positive impact across the whole sector because they benefit our partners and vendors as well as our internal programs.

Continuous technology development is critical to addressing the challenges of genetic medicines, now and in the future. ElevateBio keeps ahead on this curve, evaluating whether emerging technologies, inside and outside biotech, could fit into and advance our capabilities – and our team of super-smart thinkers bring a unique perspective to these evaluations. We hope our work to pioneer and push technology development for genetic medicines will lift the entire industry and ultimately serve more patients better.

Table 1. Examples of Novel and Emerging Technologies Evaluated by ElevateBio

Technology TypeProcess Unit OperationPurpose of Technology
Bioreactor PlatformCell Expansion & HarvestFully controlled, flexible, and automated smart cell processing platform with in-line analytics
Activation, Transduction & ExpansionCondition T cells through activation, transduction and/or expansion in incubator to increase their function in TME
Cell Sorter DeviceCell IsolationAlternative device for isolating rare blood cell populations at high purity to reduce operation time and complexity
Cell Processing DeviceStarting Material ProcessingRapid, microfluidics-based closed-system process to separate PBMCs from apheresis or whole blood
Cell Processing PlatformCell Wash & Concentrate, Cell Separation Harvest, Fill/FinishAutomated closed system for multiple unit operations to streamline manufacturing and reduce risk
Gene Modification Delivery SystemGene ModificationClosed and automated system for gentle, multiplex delivery of genetic material to cells
Lipid Nanoparticles (LNPs)Genetic Material DeliveryNon-viral cell targeted formulation for delivery of genetic material
Non-DMSO Cryoprotectant SolutionFill/FinishNatural organic compounds to protect the integrity of cells during cryopreservation
DNA Template for HDRNon-viral Genetic ModificationAlternative CRISPR templates for gene editing
Selection/Activation Reagents and cell selection kitCell Selection & ActivationAlternative reagents (including non-magnetic, nanobeads, etc.) to reduce cell process complexity and shorten vein-to-vein time, alternative kit and reagents to increase efficiency and effective phenotype and to reduce raw material manufacturing COGS
Bioreactor Harvest DeviceViral Vector Harvest UnitContinuous manufacturing, improving harvesting yields
Bioreactor Perfusion DeviceViral Vector ProductionUpstream process intensification, continuous manufacturing, improving harvesting yields
Chromatography ResinsViral Vector PurificationImprove impurity removal during viral vector downstream processing
Purification DeviceNanofiber material for vector concentration and purification
Purification ReagentChromatography-free tagging system for viral vector purification
NucleasesViral Vector Nucleic Acid DigestionImprove nucleic acid digestion during viral vector downstream processing
COGS – cost of goods; DMSO – dimethylsulfoxide; HDR – homology-direct repair; PBMCs – peripheral blood monocytes; TME – tumor microenvironment

As of September 2024

  • Page 1
  • Page 2
  • Next

Footer

ElevateBio

Investors & Media

LinkedIn Twitter

© 2026 ElevateBio. All Rights Reserved.

Privacy & Cookie Policy

  • Manufacturing & Discovery Services
  • Technologies
  • Advanced Therapies
  • Partner With Us
  • About
  • Presentations & Publications
  • Elevated Insights
  • Careers
  • Facilities
  • Contact
MENU
  • Manufacturing & Discovery Services
    • Manufacturing & Discovery Services Overview
    • Gene Editing Design & Optimization
    • Process Development
    • Analytical Development
    • cGMP Manufacturing
    • Next-Generation Sequencing
    • Quality
    • Regulatory CMC
    • Certifications & Compliance
    • Generative AI
  • Technologies
    • Technologies Overview
    • Gene Editing
    • Viral and Non-Viral Delivery
    • Messenger RNA (mRNA)
    • Induced Pluripotent Stem Cells (iPSCs)
  • Advanced Therapies
  • Partner With Us
  • About
    • About Overview
    • Our Journey
    • Our Team
    • Culture of Expedition
    • Careers
    • Investors
  • Resources
    • Resources Overview
    • Elevated Insights
    • Presentations & Publications
    • Whitepapers
  • Media
    • Media Overview
    • Press Releases
    • Featured News
  • Facilities
  • Careers
  • Contact